Tiny imperfections in diamond could offer new ways to store and read quantum information. The challenge is deciding which defects deserve the time and expense of experimental testing. A computational study now brings tools from molecular chemistry to that search, identifying promising boron- and oxygen-containing candidates.
John Mark P. Martirez, a staff research scientist at the U.S. Department of Energy’s Princeton Plasma Physics Laboratory, reports the work in ACS Central Science. His approach concentrates demanding calculations on a small group of atoms surrounding a defect. Crucially, the model still accounts for the influence of the diamond crystal beyond that group.
The results describe predicted electronic and optical properties, rather than newly demonstrated quantum computers or sensors. They identify candidates with useful characteristics and explain how their behavior relates to their chemistry. Whether those candidates can preserve quantum information reliably remains among the questions that further research must answer.
A qubit is a basic unit of quantum information, with behavior governed by quantum states rather than ordinary binary switching alone. Some diamond defects provide electronic states that researchers can manipulate with magnetic fields, light and microwaves. Their response to those inputs can also reveal information about their surroundings, making them useful for sensing.
Diamond offers an attractive host because certain defects can operate at room temperature. Its crystal structure also provides an environment in which localized defect states can be distinguished from the surrounding material. That does not mean every diamond defect becomes a useful qubit, or that room-temperature operation guarantees a complete working quantum computer.
One extensively studied example is the negatively charged nitrogen-vacancy center. A nitrogen atom replaces a carbon atom beside an empty carbon site, creating a particular arrangement of electrons. Researchers can manipulate its spin states with microwaves and obtain information through changes in fluorescence following laser excitation.
Martirez examined that familiar center alongside isolated carbon vacancies and vacancy complexes containing boron or oxygen. He considered different electrical charges and spin states, because the same atomic arrangement can behave differently when electrons are added or removed. The comparison links potential performance to both chemical composition and electronic configuration.
Accurate simulations become expensive when they must describe interacting electrons across a large piece of crystal. Defect electrons can occupy several competing configurations, making some standard approximations inadequate for detailed predictions. Molecular quantum chemistry offers more sophisticated methods, but applying them directly to large crystalline systems is difficult.
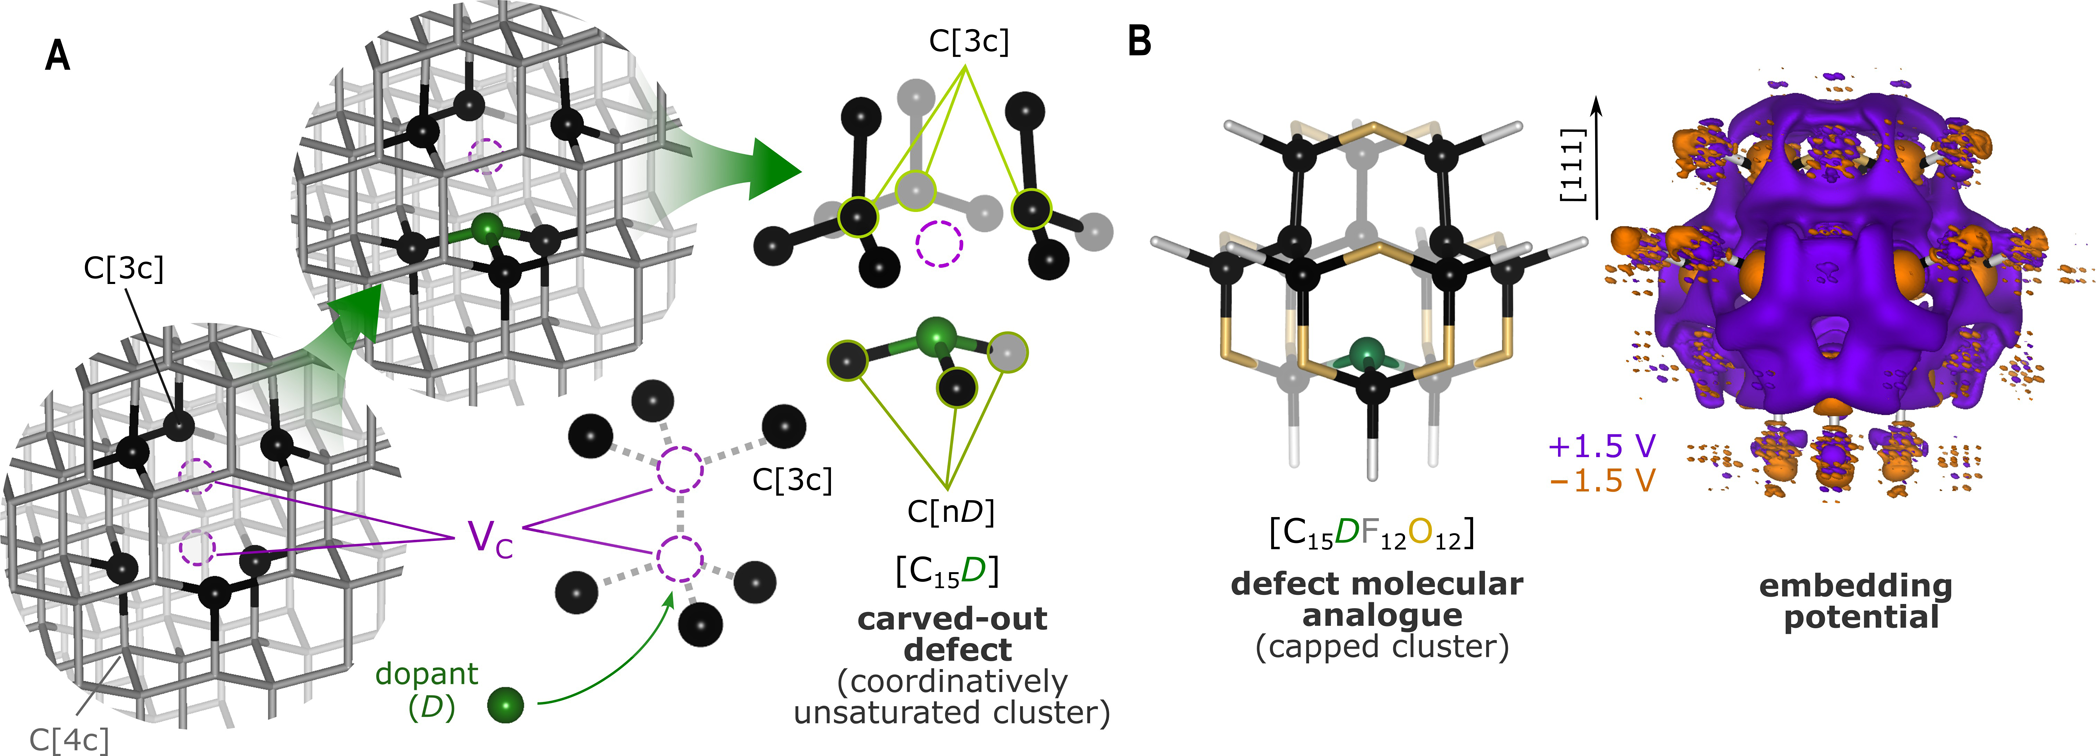
Martirez addressed that mismatch through an embedded molecular model. In one construction, the central fragment contains 15 carbon atoms and a dopant atom surrounding the vacancy. Additional fluorine and oxygen atoms cap the cut bonds in the computational fragment, producing a 40-atom model rather than a physically extracted molecule.
Those caps help make the fragment suitable for molecular calculations, but they are not the whole solution. An optimized mathematical potential represents effects of the remaining diamond, including its interaction with the defect. The method therefore reduces the expensive part of the calculation without simply pretending the surrounding crystal has disappeared.
“Because we’re treating the qubits as molecules, the cost of calculating these defects is equivalent to the cost of just modeling a simple molecule,” Martirez said. The framework combines a less expensive description of the crystalline environment with higher-level calculations of the defect itself. This makes detailed comparisons across candidate centers more tractable.
The study calculates electronic energy levels, transition strengths and natural excited-state emission lifetimes. These quantities help indicate what light a defect might absorb or emit and how readily those optical transitions occur. Molecular orbital analysis also explains how nearby carbon atoms contribute to the states often associated with the dopant.
Familiar diamond centers provide a way to check the framework against measurements. For the negatively charged nitrogen-vacancy center, calculated excitation energies agreed with experimental values to within about 0.1 electronvolt. Reproducing known properties lends support to using the approach for candidates whose experimental behavior remains less well characterized.
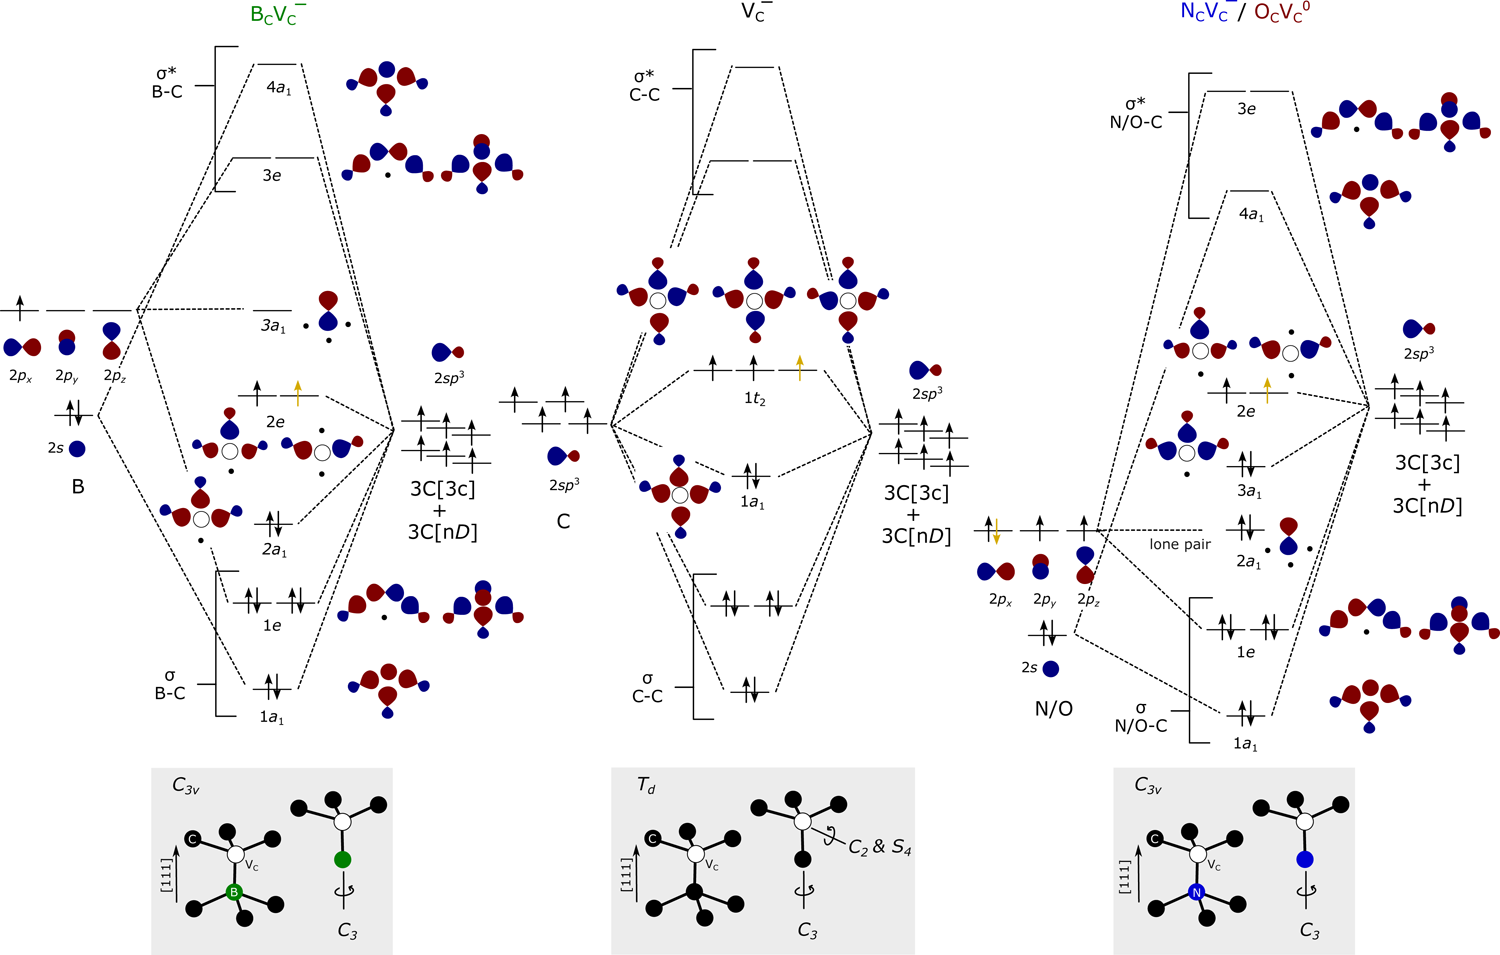
The most promising alternatives highlighted were a negatively charged boron-vacancy complex and a neutral oxygen-vacancy complex. Both have electronic structures with features resembling the established nitrogen-vacancy center. Their predicted optical transitions and additional accessible spin states make them worth examining as possible qubits.
Other charge states looked less favorable for certain applications. Some were predicted to emit weakly or lack suitable transitions between spin states for convenient optical operation. Screening can therefore help narrow the search, while explaining why an apparently small chemical change might alter a defect’s usefulness.
The oxygen-containing candidate’s predicted optical response lies at higher energies than that of the familiar nitrogen-vacancy center. The boron-containing candidate also offers a stronger transition at higher energy than its dimmer, lower-energy alternative. These differences suggest routes to defects addressed with different colors of light. The calculations place relevant oxygen transitions in the blue region and the brighter boron transition in the ultraviolet.
That could eventually help distinguish multiple kinds of qubits within one device. “One could imagine, then, combining different qubits of different responses, so you can basically probe one qubit but not the other,” Martirez said. The study proposes useful ingredients for that possibility, rather than demonstrating selective control in a working mixed-qubit system.
Brightness and timing also matter when information is read optically. Several candidate excited states had shorter calculated radiative lifetimes than the nitrogen-vacancy reference, potentially allowing more frequent signals. Those emission lifetimes describe optical relaxation, however, and must not be confused with coherence times, which measure how long useful quantum relationships persist.
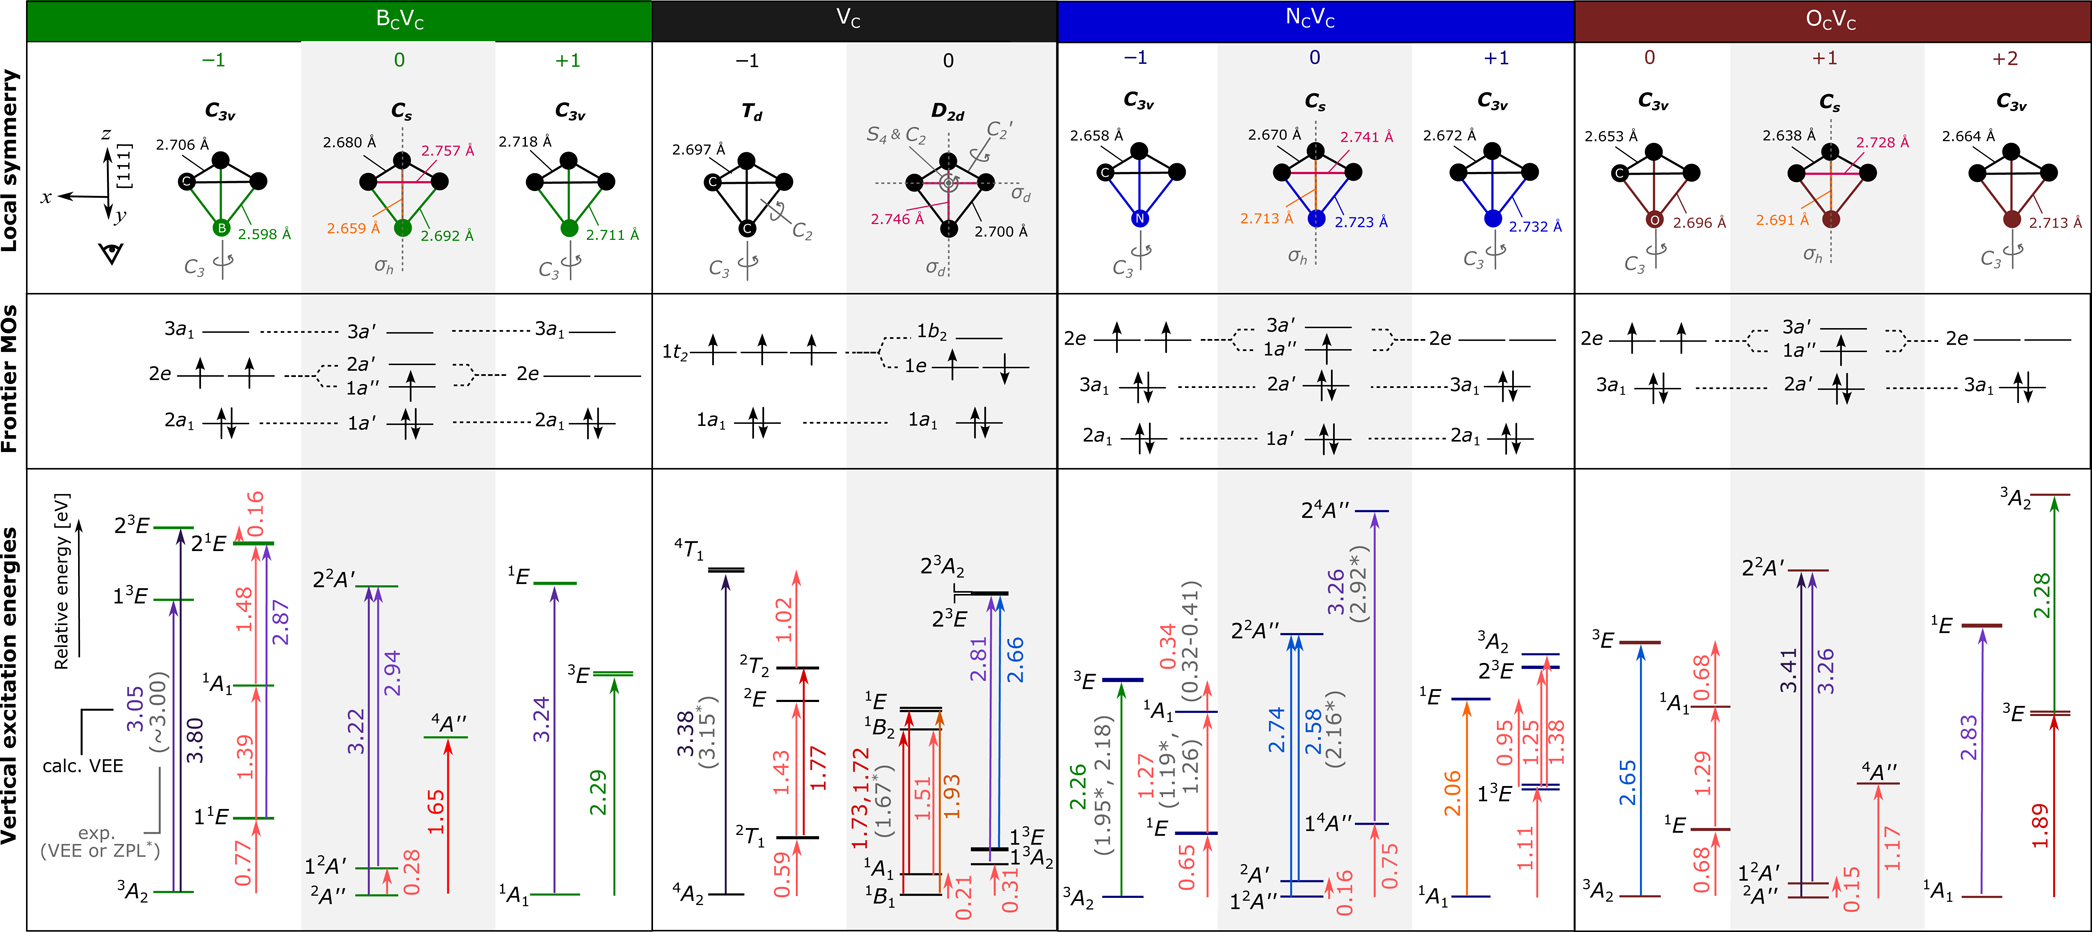
A separate stability issue complicates the oxygen candidate. The paper notes earlier calculations suggesting it can favor a different structure involving an additional vacancy. A desirable electronic response therefore does not, by itself, establish that a proposed center will be straightforward to create and maintain.
The paper explicitly lists properties still needed for a full assessment. These include interactions between electrons and crystal vibrations, spin-level splitting, transitions between spin states and coherence lifetimes. Environmental magnetic and electric fields can also affect whether a candidate remains useful outside an idealized calculation. The calculations can guide those tests, but they do not replace experimental evidence that the defects work as qubits.
The work illustrates how chemistry can guide the search through possible defect structures. Rather than treating the dopant alone as the source of useful behavior, the analysis examines the electronic system formed with neighboring atoms. That perspective gives researchers chemical explanations for differences that experiments would otherwise need to explore more broadly.
Martirez also sees a possible longer-term connection back to chemistry. Better quantum devices could eventually improve calculations of molecules and materials, extending the tools used to design them. For the candidates identified here, the immediate task is more specific: establish whether favorable calculated optical properties can be combined with stable, controllable quantum states.
These resources explain the computational framework, independent assessments and experimental benchmarks behind diamond-based quantum research.
Solid-State Point Defects through a Molecular Lens
This expert commentary explains the embedded-cluster approach and places the search beyond nitrogen-vacancy centers in context. (ACS Central Science, September 2026)
Optical properties of a diamond NV color center from capped embedded multiconfigurational correlated wavefunction theory
An earlier study benchmarks the capped embedding framework against the measured excitation energies of a well-characterized diamond defect. (npj Computational Materials, February 2026)
Accurate and convergent energetics of color centers by wavefunction theory
This independent study examines molecular cluster calculations as a way to reproduce the electronic properties of diamond color centers. (npj Computational Materials, November 2025)
Projector-Free Capped-Fragment Scheme within Density Functional Embedding Theory for Covalent and Ionic Compounds
This methodological paper introduces the capped-fragment embedding approach used to connect high-level local calculations with a larger environment. (Journal of Chemical Theory and Computation, June 2021)
Room-Temperature Quantum Bit Memory Exceeding One Second
An experimental study demonstrates long-lived quantum coherence in a carbon nuclear spin near a nitrogen-vacancy center in diamond. (Science, June 2012)
Research findings are available online in the journal ACS Central Science.
The original story “Scientists can now predict qubit behavior faster and more accurately” is published in The Brighter Side of News.
Like these kind of feel good stories? Get The Brighter Side of News’ newsletter.
The post Scientists can now predict qubit behavior faster and more accurately appeared first on The Brighter Side of News.


Leave a comment
You must be logged in to post a comment.